1
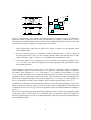
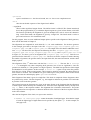
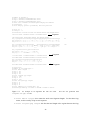
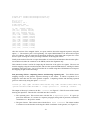
ments in each genome. In this example, these are 180, 177, and 181 in the first, second, and
third genomes, respectively.
• Total unaligned gap length: this lists the total length of the unaligned gaps.
• Total identity in aligned gaps: this lists how many characters are identical in the
aligned gaps.
• Percentage identity of chain: this lists the ratio between the total identical characters (in fragments and in aligned gaps) and the chain length in each genome. In this example,
the percentage identities are ≈ 0.66, ≈ 0.67, and ≈ 0.66 in the first, second, and third genomes,
respectively.
5.6 The synteny parameters, and the program chainer2permutation.x
Two similar regions are considered syntenic, if (1) they appear in the same order in all given genomes,
and (2) they are not interrupted by any other segment.
The line “SYNTENY=” in the parameter file specifies the parameters passed to the program chainer2permutation.x,
which performs the following tasks over the computed chains, or the filtered chains (the alignment of
these chains has nucleotide identity larger than a user-defined threshold):
1. It filters repeated sequences.
2. It computes syntenic regions and reports permutations for these segments.
3. It plots the syntenic regions w.r.t. all pairwise projections.
A fragment is considered as a repeated one, if it is overlapping with another fragment with a factor τ
of its lengt, where τ is larger than a user-defined threshold.
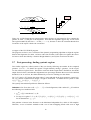
We compute the syntenic regions by applying one-dimensional chaining iteratively w.r.t. each genome.
We allow that the fragments of the chain can overlap with a factor µ of its length, where µ is larger
than a user-defined threshold.
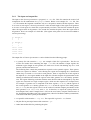
-overlap1 τ
The option -overlap1 τ permits any fragment in a chain to overlap with the next fragment
with percentage τ of its length.
-overlap2
This option allows any fragment in a chain to overlap with the next fragment with at most ℓ − 1
bp, where ℓ is the minimum region length.
-filterrep ration
The option -filterrep τ filters out any fragment overlapping with another fragment with
more than 100 × τ , 0 ≤ τ ≤ 1, of its length in any genome as a repeated one.
If no option is given, the default is not to filter repeats and allow no overlapping between the fragments
of the chain.
44